P11166
Gene name |
SLC2A1 |
Protein name |
Solute carrier family 2, facilitated glucose transporter member 1 |
Names |
Glucose transporter type 1, erythrocyte/brain, GLUT-1, HepG2 glucose transporter |
Species |
Homo sapiens (Human) |
KEGG Pathway |
hsa:6513 |
EC number |
|
Protein Class |
|
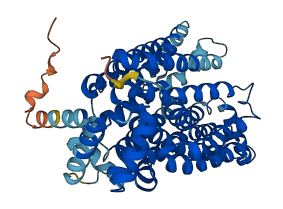
Descriptions
The autoinhibited protein was predicted that may have potential autoinhibitory elements via cis-regPred.
Autoinhibitory domains (AIDs)
Target domain |
|
Relief mechanism |
|
Assay |
cis-regPred |
Accessory elements
No accessory elements

512 variants for P11166
| Variant ID(s) | Position | Change | Description | Diseaes Association | Provenance |
|---|---|---|---|---|---|
|
RCV001249305 rs1553157935 RCV000578988 RCV002529042 |
1 | M>T | GLUT1 deficiency syndrome 1, autosomal recessive GLUT1 deficiency syndrome [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001731941 rs1570601100 RCV000812775 CA339964905 |
9 | T>M | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1333609390 RCV003141711 CA10590134 RCV000707535 |
11 | R>C | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
rs1570601060 RCV000986293 |
16 | V>missing | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
CA339964867 RCV000648083 rs1553156841 |
16 | V>L | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
CA339964861 RCV000502046 rs1345986424 |
17 | G>* | GLUT1 deficiency syndrome [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
CA339964860 rs1553156840 RCV000531748 |
17 | G>E | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000733447 rs1345986424 CA339964863 RCV001855780 |
17 | G>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
rs1557651202 CA339964846 RCV000694163 |
19 | A>V | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1570601051 RCV000820367 |
20 | V>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1165759782 RCV000820791 CA339964816 RCV001507439 |
25 | Q>K | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
RCV000705848 CA339964771 rs1557651185 |
31 | G>E | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000147518 CA018969 rs587784390 |
34 | N>D | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000030921 VAR_054755 CA018991 rs80359812 |
34 | N>I | Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS2 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV000824820 CA339964750 rs1570601007 |
34 | N>K | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
VAR_054756 RCV000995645 CA339964752 RCV001268517 RCV000819798 rs80359812 RCV002501134 |
34 | N>S | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Hereditary cryohydrocytosis with reduced stomatin GLUT1DS1; 55% of wild-type glucose uptake activity [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
| VAR_065206 | 34 | N>Y | GLUT1DS1 [UniProt] | Yes | UniProt |
|
rs796053245 RCV002517016 CA318417 RCV000189347 |
35 | A>S | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP gnomAD |
|
CA339964740 RCV000850198 rs1570600997 |
36 | P>L | Intellectual disability [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000189348 RCV001096623 CA318419 RCV000819050 rs748082803 RCV001098347 |
42 | E>A | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
CA803607 RCV001300895 rs375881934 |
44 | Y>H | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ESP ExAC TOPMed dbSNP gnomAD |
|
rs1249135503 RCV001228286 CA339962559 |
45 | N>D | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
RCV000705361 rs754791604 RCV001759415 CA803606 |
46 | Q>E | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV001711485 RCV001096621 RCV001096622 CA318421 RCV000865433 rs149998596 |
46 | Q>H | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 (dyt9) Dystonia 9 [ClinVar, Ensembl] | Yes |
ClinGen ClinVar ESP ExAC TOPMed dbSNP gnomAD |
|
RCV001226075 rs1643485738 |
47 | T>A | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
CA10581792 rs878854904 RCV000228194 |
49 | V>F | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
RCV002504382 RCV001322048 RCV001266485 COSM1342756 CA803603 rs201815571 VAR_076226 |
51 | R>H | Variant assessed as Somatic; 0.0 impact. GLUT1 deficiency syndrome 1, autosomal recessive large_intestine Hereditary cryohydrocytosis with reduced stomatin Inborn genetic diseases EIG12; unknown pathological significance [NCI-TCGA, ClinVar, Cosmic, UniProt] | Yes |
ClinGen cosmic curated ClinVar UniProt ExAC NCI-TCGA TOPMed dbSNP gnomAD |
|
CA339962418 rs201815571 RCV000533560 |
51 | R>P | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
rs796053246 RCV000189350 CA318423 RCV001038415 |
53 | G>V | GLUT1 deficiency syndrome 1, autosomal recessive Variant assessed as Somatic; impact. [ClinVar, NCI-TCGA] | Yes |
ClinGen ClinVar Ensembl NCI-TCGA dbSNP |
|
CA803602 RCV001096619 RCV002315969 RCV001096620 RCV000648090 rs765479065 |
58 | P>S | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
RCV000476048 RCV000593978 CA803599 VAR_076227 COSM236574 rs142986731 |
60 | T>M | GLUT1 deficiency syndrome 1, autosomal recessive autonomic_ganglia EIG12; unknown pathological significance; decreased glucose transport [ClinVar, Cosmic, UniProt] | Yes |
ClinGen cosmic curated ClinVar UniProt ESP ExAC TOPMed dbSNP gnomAD |
|
RCV001316126 rs1643485004 |
62 | T>I | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV002523299 RCV000763915 rs200828053 RCV000476696 RCV001580496 CA803597 |
63 | T>M | GLUT1 deficiency syndrome 1, autosomal recessive Hereditary cryohydrocytosis with reduced stomatin Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
CA21252454 VAR_013283 rs80359813 |
66 | S>F | GLUT1DS1 [UniProt] | Yes |
ClinGen UniProt Ensembl dbSNP |
|
RCV001759518 RCV000799246 CA339962024 rs1570593865 |
68 | S>L | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV001046931 rs1643484773 |
69 | V>M | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001270785 rs1643484697 |
73 | S>F | Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1553156199 RCV000539168 |
77 | M>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001246954 rs1643484447 |
77 | M>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1187210267 CA339961831 VAR_076228 |
77 | M>T | EIG12; decreased glucose transport [UniProt] | Yes |
ClinGen UniProt dbSNP gnomAD |
|
rs1643484465 RCV001204666 |
79 | G>V | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001266277 rs1643484307 |
86 | F>missing | Inborn genetic diseases [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1643484242 RCV001218614 |
87 | V>I | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs80359814 RCV000017490 CA019112 VAR_013182 |
91 | G>D | Encephalopathy due to GLUT1 deficiency GLUT1DS1; significantly decreases the transport of 3-O-methyl-D-glucose [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV000986292 rs1570593820 |
92 | R>missing | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
rs779073410 RCV001349830 |
92 | R>P | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000426262 rs202060209 VAR_069077 RCV000017499 RCV002243647 RCV000705009 RCV000850197 CA019118 RCV001843456 |
92 | R>W | Intellectual disability GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Developmental disorder Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS2 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt 1000Genomes ExAC TOPMed dbSNP gnomAD |
|
CA21252168 RCV001387742 rs80359815 RCV002249465 RCV000761655 |
93 | R>Q | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
CA019133 VAR_065207 rs267607061 RCV000648074 RCV001291641 RCV000030922 RCV000442654 RCV001548750 |
93 | R>W | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS2 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt dbSNP gnomAD |
|
CA339961448 rs1553156175 RCV000531950 |
94 | N>S | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
CA019143 rs267607060 RCV000017496 VAR_065208 |
95 | S>I | Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS2 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV001257676 rs1643481930 |
96 | M>missing | Intellectual disability [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001449666 RCV001051269 rs1643481875 |
96 | M>T | GLUT1 deficiency syndrome 1, autosomal recessive Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinVar dbSNP |
|
rs753161833 RCV000648089 RCV001532534 VAR_065209 CA803565 |
96 | M>V | GLUT1 deficiency syndrome 1, autosomal recessive Variant assessed as Somatic; 0.0002433 impact. GLUT1DS1 [ClinVar, NCI-TCGA, UniProt] | Yes |
ClinGen ClinVar UniProt ExAC NCI-TCGA dbSNP gnomAD |
|
rs1643481831 RCV001255171 |
98 | M>missing | Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000792319 COSM909705 CA339961378 rs1211054711 |
99 | M>V | GLUT1 deficiency syndrome 1, autosomal recessive endometrium [ClinVar, Cosmic] | Yes |
ClinGen cosmic curated ClinVar dbSNP gnomAD |
|
RCV000735239 rs1557646673 |
100 | N>missing | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000805606 rs1570593621 |
102 | L>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
CA803560 RCV000480070 rs76672402 RCV000662334 RCV001711886 RCV002321974 RCV000864633 RCV000311944 RCV000350448 |
104 | F>L | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 Inborn genetic diseases Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinGen ClinVar 1000Genomes ESP ExAC TOPMed dbSNP gnomAD |
|
RCV000494313 CA339961288 rs577667739 RCV002527078 |
105 | V>L | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar 1000Genomes ExAC TOPMed dbSNP gnomAD |
|
RCV000394308 RCV001721221 RCV000346798 rs577667739 COSM1667492 RCV000468529 CA318425 |
105 | V>M | GLUT1 deficiency syndrome 1, autosomal recessive large_intestine Encephalopathy due to GLUT1 deficiency Dystonia 9 [ClinVar, Cosmic] | Yes |
ClinGen cosmic curated ClinVar 1000Genomes ExAC TOPMed dbSNP gnomAD |
|
CA803556 rs74323945 RCV000730291 RCV001855633 |
108 | V>M | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
RCV001102022 RCV001100034 CA339961190 rs1399284513 |
111 | G>V | Encephalopathy due to GLUT1 deficiency Dystonia 9 [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
rs774348625 RCV001221728 CA803555 RCV002316104 |
113 | S>L | GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV001253108 rs1643480923 RCV001255340 |
114 | K>missing | Intellectual disability Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1643480570 RCV001233741 |
125 | G>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000030838 RCV000017498 RCV000030839 RCV000824987 VAR_054757 RCV000081431 CA019162 rs80359818 RCV000546969 |
126 | R>C | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 (dyt9) Dystonia 9 Hereditary cryohydrocytosis with reduced stomatin Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS1, GLUT1DS2 and DYT9; reduced transporter activity [ClinVar, Ensembl, UniProt] | Yes |
ClinGen ClinVar UniProt dbSNP gnomAD |
|
RCV002288508 RCV001387741 VAR_013183 rs80359816 RCV000081432 CA019167 RCV000017491 |
126 | R>H | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS1; significantly decreases the transport of 3-O-methyl-D-glucose and dehydroascorbic acid; 57% of wild-type glucose uptake activity [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV002271987 rs80359816 VAR_013184 RCV000017489 CA019172 |
126 | R>L | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency GLUT1DS1 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV000704712 CA803547 RCV002360818 rs751907207 |
129 | I>M | GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
rs80359819 RCV000770978 RCV001869075 CA21252051 RCV002305536 VAR_054758 |
130 | G>S | GLUT1 deficiency syndrome 1, autosomal recessive GLUT1 deficiency syndrome GLUT1DS1; 75% of wild-type glucose uptake activity [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
CA21252047 RCV003163632 RCV001214071 rs987202561 RCV002287482 |
131 | V>A | Epilepsy, idiopathic generalized, susceptibility to, 12 GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
rs1570593449 RCV000799264 |
131 | V>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001214425 rs1643480228 |
133 | C>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
CA16043369 rs1057518953 RCV000415325 RCV000458446 |
134 | G>S | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000648085 CA339960777 rs1057517822 |
140 | V>L | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1057517822 RCV000414747 CA16042396 RCV001850982 |
140 | V>M | GLUT1 deficiency syndrome 1, autosomal recessive Variant assessed as Somatic; impact. [ClinVar, NCI-TCGA] | Yes |
ClinGen ClinVar Ensembl NCI-TCGA dbSNP |
|
rs1570593475 RCV000800135 |
142 | M>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
CA21252032 VAR_013284 rs80359820 |
146 | E>K | GLUT1DS1 [UniProt] | Yes |
ClinGen UniProt Ensembl dbSNP |
|
rs1643479604 RCV001069131 |
148 | S>L | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
| VAR_076229 | 149 | P>A | EIG12; unknown pathological significance [UniProt] | Yes | UniProt |
| VAR_054759 | 153 | R>C | GLUT1DS1; 44% of wild-type glucose uptake activity [UniProt] | Yes | UniProt |
|
rs794727642 CA019189 RCV000178275 COSM3400787 VAR_065210 RCV001852209 |
153 | R>H | GLUT1 deficiency syndrome 1, autosomal recessive Variant assessed as Somatic; impact. central_nervous_system GLUT1DS2 [ClinVar, NCI-TCGA, Cosmic, UniProt] | Yes |
ClinGen cosmic curated ClinVar UniProt Ensembl NCI-TCGA dbSNP |
|
RCV001253255 rs794727642 CA318491 RCV000189394 RCV001255335 |
153 | R>L | Intellectual disability Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV002265788 CA339960546 rs794727642 RCV000520301 |
153 | R>P | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
| VAR_065211 | 155 | A>V | GLUT1DS1 [UniProt] | Yes | UniProt |
|
rs1553156157 CA339960497 RCV000648076 |
157 | G>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000415305 rs1057518821 |
158 | T>missing | Cerebellar ataxia [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001039591 CA339960460 rs1472329184 |
160 | H>Y | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
CA339960441 rs1413339367 RCV000624807 RCV001091412 RCV001389445 |
161 | Q>* | GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
CA339960438 RCV000648075 rs1553156154 |
161 | Q>P | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
VAR_065212 RCV001865327 CA16603717 RCV000429855 RCV002468576 rs1057520545 |
165 | V>I | GLUT1 deficiency syndrome 1, autosomal recessive Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS2 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV001253375 rs773339124 RCV001255358 |
167 | G>R | Intellectual disability Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001851153 rs80359832 VAR_054760 RCV000478031 RCV002470868 |
169 | L>missing | GLUT1 deficiency syndrome 1, autosomal recessive GLUT1DS1; 48% of wild-type glucose uptake activity Childhood onset GLUT1 deficiency syndrome 2 [ClinVar, UniProt] | Yes |
ClinVar UniProt dbSNP |
|
rs80359832 VAR_054760 |
169 | L>del | GLUT1DS1; 48% of wild-type glucose uptake activity [UniProt] | Yes |
UniProt dbSNP |
|
RCV001814283 rs1017082606 COSM3400784 CA21251974 RCV001215093 RCV001838656 |
171 | A>T | GLUT1 deficiency syndrome 1, autosomal recessive Variant assessed as Somatic; impact. central_nervous_system Dystonia 9 [ClinVar, NCI-TCGA, Cosmic] | Yes |
ClinGen cosmic curated ClinVar NCI-TCGA TOPMed dbSNP gnomAD |
|
RCV001046838 rs1643471482 |
174 | F>C | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1085308009 RCV000489014 RCV002298620 CA339958920 |
175 | G>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1643471410 RCV001337445 |
176 | L>P | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000815374 CA339958866 rs1570592960 |
178 | S>A | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1643471237 RCV001298353 |
182 | N>S | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1570592942 CA339958710 RCV000806967 RCV002534834 |
185 | L>P | GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
CA339958705 rs1570592933 RCV000801605 RCV000986291 |
186 | W>* | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1643471012 RCV001337709 |
186 | W>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001225131 rs1459390244 CA339958697 |
187 | P>S | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
rs878853161 RCV000224978 |
192 | I>missing | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000795314 rs746393667 CA339958670 |
192 | I>V | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV000648087 rs1553156090 |
193 | I>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001865350 rs1057521967 RCV000429681 CA16603691 |
200 | Q>* | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
CA21251449 rs1050933017 RCV001321659 RCV001028107 RCV001532532 |
201 | C>G | GLUT1 deficiency syndrome 1, autosomal recessive Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
rs779779804 RCV001304146 CA803492 |
202 | I>M | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
CA339958604 RCV001071023 rs758200582 |
203 | V>M | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
rs1643470058 RCV001299553 |
206 | F>C | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000986290 rs1570592844 |
209 | E>missing | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001205552 rs1643470035 |
209 | E>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000986289 CA339958564 rs1387203768 |
209 | E>* | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
RCV001326291 RCV003135909 rs1387203768 CA339958565 RCV001267166 |
209 | E>K | Variant assessed as Somatic; 0.0 impact. GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [NCI-TCGA, ClinVar] | Yes |
ClinGen ClinVar NCI-TCGA dbSNP gnomAD |
|
RCV000785926 rs1557646075 |
209 | E>missing | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001365001 rs796053247 RCV000189354 CA318431 |
211 | P>S | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
CA019231 RCV000814788 RCV000255028 rs387907312 VAR_065213 RCV000030840 |
212 | R>C | GLUT1 deficiency syndrome 1, autosomal recessive Dystonia 9 (dyt9) Dystonia 9 GLUT1DS1 and DYT9 [ClinVar, Ensembl, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
VAR_065214 RCV000255929 rs886039517 CA10588297 RCV001390267 RCV001253348 |
212 | R>H | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency GLUT1DS1 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
rs1570592813 RCV001664443 RCV000821549 CA339958526 |
215 | L>F | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
CA019237 RCV000690814 rs147249343 RCV000723629 |
218 | R>C | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ESP ExAC TOPMed dbSNP gnomAD |
|
RCV001098263 RCV001098262 RCV001266484 rs374080633 RCV000657887 RCV001079315 CA803486 |
218 | R>H | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ESP ExAC TOPMed dbSNP gnomAD |
| VAR_076230 | 218 | R>S | EIG12; decreased glucose transport [UniProt] | Yes | UniProt |
|
rs1557646057 RCV000707556 CA339958475 |
222 | N>K | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs397514564 VAR_065215 RCV000032904 CA019242 |
223 | R>P | Epilepsy, idiopathic generalized, susceptibility to, 12 Epilepsy, idiopathic generalized, susceptibility to, 12 (eig12) EIG12; mild phenotype; reduced transporter activity; impaired phosphorylation by PKC [ClinVar, Ensembl, UniProt] | Yes |
ClinGen ClinVar UniProt ExAC TOPMed dbSNP gnomAD |
|
RCV001546081 RCV001045125 rs397514564 CA803484 RCV002363593 VAR_076231 RCV001098259 |
223 | R>Q | GLUT1 deficiency syndrome 1, autosomal recessive Epilepsy, idiopathic generalized, susceptibility to, 12 (eig12) Dystonia 9 Inborn genetic diseases EIG12; unknown pathological significance; no effect on glucose transport; impaired phosphorylation by PKC [ClinVar, Ensembl, UniProt] | Yes |
ClinGen ClinVar UniProt ExAC TOPMed dbSNP gnomAD |
|
RCV000546488 rs796053248 RCV000189355 RCV002288795 CA318433 VAR_065216 RCV000763337 |
223 | R>W | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Hereditary cryohydrocytosis with reduced stomatin GLUT1DS1; impaired phosphorylation by PKC [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt dbSNP gnomAD |
|
RCV001346218 rs1643467923 |
230 | K>N | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs387907313 CA019279 COSM909703 RCV000189356 RCV000030841 VAR_069078 RCV001063268 |
232 | R>C | Epilepsy, idiopathic generalized, susceptibility to, 12 Variant assessed as Somatic; 0.0 impact. GLUT1 deficiency syndrome 1, autosomal recessive large_intestine endometrium Epilepsy, idiopathic generalized, susceptibility to, 12 (eig12) EIG12; the mutant protein is expressed at the cell surface but has mildly decreased glucose uptake (70%) compared to wild-type [ClinVar, NCI-TCGA, Cosmic, Ensembl, UniProt] | Yes |
ClinGen cosmic curated ClinVar UniProt ExAC NCI-TCGA dbSNP gnomAD |
|
CA803465 RCV000761654 RCV001855704 rs139412383 COSM3689698 RCV001823005 |
232 | R>H | Epilepsy, idiopathic generalized, susceptibility to, 12 Variant assessed as Somatic; 0.0 impact. GLUT1 deficiency syndrome 1, autosomal recessive large_intestine [ClinVar, NCI-TCGA, Cosmic] | Yes |
ClinGen cosmic curated ClinVar ESP ExAC NCI-TCGA TOPMed dbSNP gnomAD |
|
RCV000545154 CA803463 rs375853334 RCV002476181 |
233 | G>R | GLUT1 deficiency syndrome 1, autosomal recessive Hereditary cryohydrocytosis with reduced stomatin [ClinVar] | Yes |
ClinGen ClinVar ESP ExAC TOPMed dbSNP gnomAD |
|
RCV000224228 rs794729221 RCV001333547 RCV000184042 CA019289 |
242 | Q>* | Intellectual disability Encephalopathy due to GLUT1 deficiency Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
| VAR_076232 | 243 | E>V | EIG12; decreased glucose transport [UniProt] | Yes | UniProt |
|
RCV000578223 rs1553156069 |
244 | M>missing | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001251646 rs1064795363 |
244 | M>T | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1570592604 RCV001542521 RCV001003570 RCV001049017 |
246 | E>missing | Global developmental delay GLUT1 deficiency syndrome 1, autosomal recessive Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001067060 RCV002390305 RCV001507438 RCV000147530 CA019296 rs587784395 |
249 | R>Q | Variant assessed as Somatic; 0.0 impact. GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [NCI-TCGA, ClinVar] | Yes |
ClinGen ClinVar ExAC NCI-TCGA TOPMed dbSNP gnomAD |
|
RCV001950657 rs796053250 CA318436 |
249 | R>W | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000147531 CA019302 rs587784396 RCV000189359 |
250 | Q>* | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1643466842 RCV001337899 |
254 | E>K | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001096518 RCV001704070 RCV000147532 RCV002390306 RCV000805096 RCV001096519 rs5811 CA019308 |
255 | K>T | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
CA21251080 VAR_013185 rs121909738 |
256 | K>E | GLUT1DS1 [UniProt] | Yes |
ClinGen UniProt TOPMed dbSNP gnomAD |
|
CA019315 RCV000017488 RCV000189388 rs80359822 RCV002271986 |
256 | K>V | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV001326528 RCV000993977 rs1557645931 |
256 | K>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000704304 CA339957776 rs1557645921 |
260 | L>P | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1302959508 CA339957718 RCV003128772 RCV001281367 |
263 | F>L | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
CA019324 RCV002317055 rs766376173 RCV000467294 RCV000179465 |
264 | R>C | GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
RCV000585350 rs758919432 CA803454 RCV001041007 |
264 | R>H | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV000226026 CA803453 rs574211861 |
266 | P>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV000805007 rs199639208 CA803451 RCV002261220 |
267 | A>T | Variant assessed as Somatic; 0.0 impact. GLUT1 deficiency syndrome 1, autosomal recessive [NCI-TCGA, ClinVar] | Yes |
ClinGen ClinVar 1000Genomes ESP ExAC NCI-TCGA TOPMed dbSNP gnomAD |
|
rs1553156058 RCV000648084 CA339957589 |
267 | A>V | GLUT1 deficiency syndrome 1, autosomal recessive Variant assessed as Somatic; impact. [ClinVar, NCI-TCGA] | Yes |
ClinGen ClinVar Ensembl NCI-TCGA dbSNP |
|
RCV001450120 CA318493 RCV001532531 rs200247956 |
269 | R>C | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ESP ExAC TOPMed dbSNP gnomAD |
|
rs1411827478 RCV001332779 RCV000823780 CA339957541 |
269 | R>H | GLUT1 deficiency syndrome 1, autosomal recessive Dystonia 9 [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
RCV001853647 RCV000624470 rs1553156053 RCV000521391 CA339957526 |
270 | Q>* | GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1557645879 CA339957479 RCV001244532 |
271 | P>L | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV001232005 rs1643465735 |
272 | I>L | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000147534 RCV001851890 VAR_054761 CA019335 rs121909740 RCV000189361 RCV000017494 |
275 | A>T | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS2; the mutation decreases glucose transport but does not affect cation permeability [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt dbSNP gnomAD |
|
RCV000559813 CA339957303 rs1553156051 RCV003153721 |
279 | Q>* | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000498692 rs1553156047 |
282 | Q>missing | Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinVar dbSNP |
|
CA16603718 RCV000432773 rs1057521066 RCV002524897 RCV002274028 |
282 | Q>* | Epilepsy, idiopathic generalized, susceptibility to, 12 GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
| VAR_054762 | 282 | Q>del | GLUT1DS2; accompanied by hemolytic anemia and altered erythrocyte ion concentrations; the mutation decreases glucose transport and causes a cation leak that alteres intracellular concentrations of sodium potassium and calcium [UniProt] | Yes | UniProt |
|
rs587784397 RCV000147535 CA019340 RCV003137647 |
283 | Q>* | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
VAR_076233 RCV000202596 CA213395 RCV003221858 rs864309514 |
286 | G>D | Hereditary cryohydrocytosis with reduced stomatin SDCHCN; no effect on protein abundance; no effect on localization to the plasma membrane; loss of D-glucose transporter activity; increased cation leakage [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
CA339957167 RCV000850196 rs1431778557 RCV001003569 |
287 | I>L | Intellectual disability Microcephaly [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
rs1643465264 RCV001297161 |
288 | N>I | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001205930 rs1643461834 |
291 | F>S | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
| VAR_069079 | 292 | Y>YY | GLUT1DS1 [UniProt] | Yes | UniProt |
|
rs1557645723 RCV001318582 |
294 | S>C | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1557645723 CA339956925 RCV002318217 |
294 | S>F | Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
| VAR_065784 | 294 | S>P | GLUT1DS2 [UniProt] | Yes | UniProt |
|
VAR_054763 CA277218 RCV002272166 RCV000189397 RCV000193872 rs80359823 RCV001380032 |
295 | T>M | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency GLUT1 deficiency syndrome GLUT1DS1; 75% of wild-type glucose uptake activity [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV000363287 RCV001721222 RCV000463634 RCV000306227 rs148518827 CA318444 RCV002317646 |
299 | E>K | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar 1000Genomes ESP ExAC TOPMed dbSNP gnomAD |
|
RCV001771998 RCV000703738 CA339956790 rs1425773776 |
301 | A>G | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP gnomAD |
|
rs1425773776 CA339956787 RCV001305693 RCV001779153 |
301 | A>V | GLUT1 deficiency syndrome 1, autosomal recessive Variant assessed as Somatic; impact. Encephalopathy due to GLUT1 deficiency [ClinVar, NCI-TCGA] | Yes |
ClinGen ClinVar NCI-TCGA TOPMed dbSNP gnomAD |
|
CA339956762 RCV001351725 rs1205631854 VAR_065217 RCV000761653 |
303 | V>L | GLUT1 deficiency syndrome 1, autosomal recessive found in a patient with GLUT1 deficiency syndrome [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt TOPMed dbSNP gnomAD |
|
rs80359824 VAR_013285 CA21250649 RCV001089897 |
310 | T>I | Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS1 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
CA211935 rs794727870 RCV000209836 |
313 | S>F | Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000281833 rs886041590 RCV001855067 |
314 | G>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs121909739 CA019364 RCV000153967 RCV000473987 RCV002288509 RCV001253635 RCV000017493 VAR_054764 |
314 | G>S | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS2; the mutation decreases glucose transport but does not affect cation permeability [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
CA318446 rs780528770 RCV000189365 RCV002444767 RCV001064748 |
315 | I>M | GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
rs920394412 RCV001037132 CA21250594 RCV000585123 |
316 | V>I | GLUT1 deficiency syndrome 1, autosomal recessive Variant assessed as Somatic; impact. [ClinVar, NCI-TCGA] | Yes |
ClinGen ClinVar NCI-TCGA TOPMed dbSNP |
| VAR_065218 | 317 | N>T | GLUT1DS2 [UniProt] | Yes | UniProt |
|
rs746091725 RCV000513651 RCV001230574 CA803400 |
318 | T>M | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
rs1060501354 RCV000470279 CA16610098 |
322 | V>A | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1643460170 RCV001325248 |
323 | V>A | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
CA803398 COSM3419240 RCV002379309 RCV000791499 RCV000439467 rs749426767 |
323 | V>M | Variant assessed as Somatic; 0.0 impact. GLUT1 deficiency syndrome 1, autosomal recessive large_intestine Inborn genetic diseases [NCI-TCGA, ClinVar, Cosmic] | Yes |
ClinGen cosmic curated ClinVar ExAC NCI-TCGA TOPMed dbSNP gnomAD |
|
VAR_065219 RCV000458906 CA318448 RCV001781556 rs796053253 RCV000189366 |
324 | S>L | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency GLUT1DS2; mild phenotype; reduced transporter activity [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
| VAR_065220 | 329 | E>Q | GLUT1DS1; stabilizes the inward-open conformation [UniProt] | Yes | UniProt |
|
rs1040371265 RCV001223859 CA21250211 |
331 | A>T | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP gnomAD |
|
rs1643457707 RCV001221345 |
332 | G>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000792856 VAR_065221 CA339956017 RCV001814181 rs1553155986 RCV001091411 RCV000517267 RCV002289706 |
333 | R>Q | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS1 and GLUT1DS2 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV002271990 CA019390 RCV001390266 RCV002312729 VAR_013286 RCV000763336 RCV000180300 rs80359825 RCV000500584 RCV002288787 |
333 | R>W | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Hereditary cryohydrocytosis with reduced stomatin Inborn genetic diseases GLUT1 deficiency syndrome Childhood onset GLUT1 deficiency syndrome 2 GLUT1DS1; 43% of wild-type glucose uptake activity [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV000802557 CA21250202 RCV001507437 rs892715050 RCV000624225 |
334 | R>Q | GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
RCV000147517 rs587784389 |
336 | L>missing | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000660366 RCV000180302 CA018956 rs755946345 |
336 | L>V | Epilepsy, idiopathic generalized, susceptibility to, 12 [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
CA339955933 rs1553155982 RCV000655968 |
337 | H>L | Childhood epilepsy with centrotemporal spikes [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000189370 CA018983 RCV002317061 RCV000724044 rs141619735 RCV000768319 RCV001781538 RCV000515255 RCV001079673 |
339 | I>T | Epilepsy, idiopathic generalized, susceptibility to, 12 GLUT1 deficiency syndrome 1, autosomal recessive Hereditary cryohydrocytosis with reduced stomatin Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ESP ExAC TOPMed dbSNP gnomAD |
|
RCV000828444 RCV001858425 rs763241827 CA803367 |
342 | A>T | Variant assessed as Somatic; 0.0 impact. GLUT1 deficiency syndrome 1, autosomal recessive [NCI-TCGA, ClinVar] | Yes |
ClinGen ClinVar ExAC NCI-TCGA TOPMed dbSNP gnomAD |
|
rs1643457017 RCV001249689 |
344 | M>missing | Hereditary cryohydrocytosis with reduced stomatin [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000585705 rs1553155973 |
345 | A>missing | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
rs769943554 RCV000316012 RCV000648088 RCV002390506 RCV000372948 RCV000192772 CA205824 RCV000725496 |
345 | A>V | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Dystonia 9 Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
rs1643456790 RCV001321509 |
348 | A>T | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001060599 rs916926401 CA21250142 |
350 | L>V | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000509454 CA803361 RCV001764508 rs780529723 RCV001217353 |
354 | A>T | GLUT1 deficiency syndrome 1, autosomal recessive Dystonia 9 [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
rs1643456257 RCV001044468 |
355 | L>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000147519 rs587784391 |
362 | P>* | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000804759 rs1570591031 CA339954420 |
363 | W>* | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
CA21249258 RCV000795449 rs976075779 |
366 | Y>C | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
CA318461 RCV000189374 RCV003114343 rs796053259 |
370 | V>A | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
rs751573593 RCV001049146 CA318459 RCV001705042 COSM3386171 RCV002252037 |
370 | V>M | GLUT1 deficiency syndrome 1, autosomal recessive pancreas [ClinVar, Cosmic] | Yes |
ClinGen cosmic curated ClinVar ExAC TOPMed dbSNP gnomAD |
|
rs1338947760 RCV000690859 CA339954246 |
371 | A>V | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
RCV001240060 rs1643443081 |
372 | I>N | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV003147392 rs796053260 RCV000189375 CA318463 |
376 | V>M | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
| VAR_065222 | 382 | G>D | GLUT1DS1 [UniProt] | Yes | UniProt |
|
RCV001333545 rs1643442833 |
383 | P>S | Dystonia 9 [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000624680 CA339953989 rs1553155891 |
385 | P>L | Variant assessed as Somatic; impact. Inborn genetic diseases [NCI-TCGA, ClinVar] | Yes |
ClinGen ClinVar Ensembl NCI-TCGA dbSNP |
|
CA318467 rs796053262 RCV000648080 |
386 | I>S | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV001855651 rs764168088 RCV000731093 CA803330 RCV001265826 |
391 | V>M | GLUT1 deficiency syndrome 1, autosomal recessive Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
RCV000648091 rs1553155887 |
393 | E>missing | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1643442142 RCV001309039 |
399 | P>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000463311 rs796053263 RCV001267280 RCV000189378 CA318469 RCV003137767 |
400 | R>C | GLUT1 deficiency syndrome 1, autosomal recessive Dystonia 9 Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV002347986 CA10602778 RCV000678305 RCV000364606 RCV000694332 rs776095655 |
400 | R>H | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
rs776095655 RCV000648086 CA803328 |
400 | R>L | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV000995644 rs1570590905 |
400 | R>S | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinVar dbSNP |
|
rs1643441930 RCV001267666 |
401 | P>L | SLC2A1-Related Disorders [ClinVar] | Yes |
ClinVar dbSNP |
| VAR_065223 | 405 | A>D | GLUT1DS1 [UniProt] | Yes | UniProt |
|
rs148800393 CA318495 RCV000648102 |
406 | V>I | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar 1000Genomes ESP ExAC TOPMed dbSNP gnomAD |
|
RCV002290466 rs1570590876 CA339953730 RCV000819910 |
408 | G>D | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV001224070 rs398123069 CA019025 VAR_076234 RCV001588909 RCV000082869 |
411 | N>S | Epilepsy, idiopathic generalized, susceptibility to, 12 GLUT1 deficiency syndrome 1, autosomal recessive Epilepsy, idiopathic generalized, susceptibility to, 12 (eig12) EIG12; decreased glucose transport [ClinVar, Ensembl, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV002318614 rs1557645002 CA339953697 |
412 | W>* | Inborn genetic diseases [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1570590859 RCV000995643 CA339953701 |
412 | W>G | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs144389023 CA803323 RCV000812546 |
417 | I>T | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ESP ExAC TOPMed dbSNP gnomAD |
|
RCV001218801 rs139722450 |
419 | G>D | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001267367 rs1643441134 |
421 | C>* | Inborn genetic diseases [ClinVar] | Yes |
ClinVar dbSNP |
|
CA339953639 rs1557644984 RCV000684767 |
421 | C>R | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
CA339953632 rs1570590834 RCV002283514 RCV000808808 |
422 | F>L | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs587784392 RCV001204221 |
423 | Q>P | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV002517900 RCV000189379 CA318471 rs796053264 |
424 | Y>* | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
rs1643440876 RCV001227926 |
426 | E>* | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000189399 RCV001208558 CA318497 rs796053274 |
429 | C>S | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar TOPMed dbSNP |
|
rs75485205 RCV000173823 CA019039 RCV002516600 |
432 | Y>* | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar 1000Genomes ESP ExAC TOPMed dbSNP gnomAD |
|
RCV001348127 rs1643436553 |
432 | Y>C | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001003340 CA339953490 rs1570590528 |
434 | F>V | Childhood onset GLUT1 deficiency syndrome 2 [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000648081 CA339953472 rs1407189135 |
435 | I>V | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
| VAR_076235 | 435 | I>del | SDCHCN; no effect on protein abundance; no effect on localization to the plasma membrane; loss of D-glucose transporter activity; increased cation leakage [UniProt] | Yes | UniProt |
|
RCV000622580 RCV000202573 rs864309522 RCV001280695 |
436 | I>missing | Hereditary cryohydrocytosis with reduced stomatin Inborn genetic diseases [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001233123 rs1643436127 |
446 | I>V | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
rs80359828 CA019048 RCV000017487 |
449 | Y>* | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
rs1216296247 RCV002546597 RCV001332778 |
450 | F>I | GLUT1 deficiency syndrome 1, autosomal recessive Dystonia 9 [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV000017486 CA019055 rs80359829 |
456 | K>* | Encephalopathy due to GLUT1 deficiency [ClinVar] | Yes |
ClinGen ClinVar Ensembl dbSNP |
|
RCV000082868 RCV000423069 CA019067 rs13306758 RCV000762930 VAR_076236 RCV000701550 |
458 | R>W | Epilepsy, idiopathic generalized, susceptibility to, 12 GLUT1 deficiency syndrome 1, autosomal recessive Epilepsy, idiopathic generalized, susceptibility to, 12 (eig12) Hereditary cryohydrocytosis with reduced stomatin EIG12; decreased glucose transport [ClinVar, Ensembl, UniProt] | Yes |
ClinGen ClinVar UniProt 1000Genomes ESP ExAC TOPMed dbSNP gnomAD |
|
RCV000766834 RCV001049565 RCV000189380 rs138139624 CA318473 |
466 | G>S | Variant assessed as Somatic; 0.0 impact. GLUT1 deficiency syndrome 1, autosomal recessive [NCI-TCGA, ClinVar] | Yes |
ClinGen ClinVar ESP ExAC NCI-TCGA dbSNP gnomAD |
|
RCV001548577 rs267607059 CA019072 RCV002271988 RCV000017500 VAR_069080 |
468 | R>W | GLUT1 deficiency syndrome 1, autosomal recessive Encephalopathy due to GLUT1 deficiency GLUT1DS1 [ClinVar, UniProt] | Yes |
ClinGen ClinVar UniProt Ensembl dbSNP |
|
RCV002318155 rs1264369249 |
469 | Q>missing | Inborn genetic diseases [ClinVar] | Yes |
ClinVar dbSNP |
|
CA16043367 RCV000415296 RCV003114532 RCV001770291 rs201748668 RCV001197510 |
469 | Q>H | Epilepsy, idiopathic generalized, susceptibility to, 12 GLUT1 deficiency syndrome 1, autosomal recessive Migraine [ClinVar] | Yes |
ClinGen ClinVar 1000Genomes dbSNP |
|
RCV000189381 RCV000189382 RCV000687947 CA318475 RCV000766835 CA318477 RCV000692340 rs572648977 |
470 | G>R | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar 1000Genomes ExAC TOPMed dbSNP gnomAD |
|
RCV000189383 rs572648977 CA318478 RCV001054891 |
470 | G>W | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar 1000Genomes ExAC TOPMed dbSNP gnomAD |
|
RCV001320328 rs955043564 CA21248757 |
471 | G>V | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar dbSNP gnomAD |
|
rs774241047 RCV001308097 CA803280 RCV000523437 |
473 | S>N | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV001042387 CA803279 rs770901544 |
475 | S>N | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV001037106 rs1643434594 |
479 | P>L | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinVar dbSNP |
|
RCV001351933 rs749067830 RCV001563011 CA803278 |
479 | P>T | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC TOPMed dbSNP gnomAD |
|
CA803277 rs756304012 RCV001373403 RCV000840858 |
480 | E>K | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV000648077 rs748340730 CA318499 RCV000189400 |
482 | L>P | GLUT1 deficiency syndrome 1, autosomal recessive [ClinVar] | Yes |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
RCV001862694 VAR_065224 rs1159593580 RCV001091410 |
485 | P>L | GLUT1 deficiency syndrome 1, autosomal recessive GLUT1DS1; creates a dileucine internalization motif that promotes recruitment of clathrin and mislocalization of the protein to endocytic compartments [ClinVar, UniProt] | Yes |
ClinVar dbSNP UniProt |
|
CA339965890 rs1271529267 |
2 | E>D | No |
ClinGen gnomAD |
|
|
rs1232724729 CA339965888 |
3 | P>S | No |
ClinGen gnomAD |
|
|
rs1335592731 CA339965880 |
4 | S>N | No |
ClinGen gnomAD |
|
|
rs1295646319 CA339965876 |
4 | S>R | No |
ClinGen TOPMed gnomAD |
|
|
CA339965875 rs780680200 |
5 | S>C | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA803685 rs780680200 |
5 | S>G | No |
ClinGen ExAC TOPMed gnomAD |
|
|
rs1326368803 CA339965873 |
5 | S>N | No |
ClinGen gnomAD |
|
|
rs150921182 CA803638 |
11 | R>H | No |
ClinGen ESP ExAC gnomAD |
|
|
CA803637 rs774411009 |
13 | M>V | No |
ClinGen ExAC gnomAD |
|
|
CA339964871 rs1288030829 |
15 | A>D | No |
ClinGen gnomAD |
|
|
CA339964873 rs1393626431 |
15 | A>S | No |
ClinGen TOPMed |
|
|
CA339964869 rs1553156841 |
16 | V>M | No |
ClinGen Ensembl |
|
| TCGA novel | 17 | G>A | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
rs1322852314 CA339964856 |
18 | G>* | No |
ClinGen gnomAD |
|
|
RCV000175828 CA019329 rs794727283 |
27 | G>D | No |
ClinGen ClinVar dbSNP gnomAD |
|
|
CA318489 rs796053273 |
27 | G>S | No |
ClinGen Ensembl |
|
|
rs1186274140 CA339964779 |
30 | T>A | No |
ClinGen gnomAD |
|
|
rs773791632 CA803635 |
35 | A>V | No |
ClinGen ExAC gnomAD |
|
|
rs770013950 CA803634 |
37 | Q>E | No |
ClinGen ExAC gnomAD |
|
|
rs770013950 CA339964738 |
37 | Q>K | No |
ClinGen ExAC gnomAD |
|
|
rs1467764389 CA339964729 |
38 | K>T | No |
ClinGen gnomAD |
|
|
rs1570593965 CA339962675 |
39 | V>G | No |
ClinGen Ensembl |
|
|
CA803608 rs769722007 |
41 | E>K | No |
ClinGen ExAC gnomAD |
|
|
CA803605 rs571371116 |
47 | T>I | No |
ClinGen 1000Genomes ExAC gnomAD |
|
|
rs1231935211 CA339962452 |
49 | V>A | No |
ClinGen TOPMed |
|
|
CA339962457 rs878854904 |
49 | V>I | Variant assessed as Somatic; 0.0 impact. [NCI-TCGA] | No |
ClinGen NCI-TCGA gnomAD |
|
rs1643485509 RCV001310847 |
52 | Y>N | No |
ClinVar dbSNP |
|
|
CA339962383 rs1235624366 |
53 | G>R | No |
ClinGen gnomAD |
|
|
RCV000177260 CA019088 rs794727508 |
55 | S>T | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
RCV001008312 rs1570593893 |
60 | T>missing | No |
ClinVar dbSNP |
|
|
rs142986731 RCV001091413 |
60 | T>R | No |
ClinVar dbSNP |
|
|
RCV001008490 rs1570593881 |
63 | T>missing | No |
ClinVar dbSNP |
|
|
rs1474108894 CA339962087 |
65 | W>C | No |
ClinGen TOPMed |
|
| TCGA novel | 70 | A>T | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
RCV000189385 rs796053266 |
70 | A>missing | No |
ClinVar dbSNP |
|
|
RCV000192838 CA205940 rs776583130 |
77 | M>V | No |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
|
CA803591 rs768621727 |
84 | G>S | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA803590 rs746796254 |
85 | L>R | No |
ClinGen ExAC gnomAD |
|
|
rs961569873 CA21252395 |
89 | R>G | No |
ClinGen Ensembl |
|
|
rs370031715 CA803589 |
89 | R>H | No |
ClinGen ESP ExAC TOPMed gnomAD |
|
|
rs779073410 CA803588 |
92 | R>Q | No |
ClinGen ExAC TOPMed gnomAD |
|
|
rs1053525817 CA339961387 |
98 | M>R | No |
ClinGen gnomAD |
|
|
CA21252155 rs1053525817 |
98 | M>T | No |
ClinGen gnomAD |
|
|
rs1211054711 CA339961373 |
99 | M>L | No |
ClinGen gnomAD |
|
|
rs781571701 CA803564 |
100 | N>S | No |
ClinGen ExAC gnomAD |
|
|
CA339961331 rs1451513110 |
102 | L>V | No |
ClinGen gnomAD |
|
|
rs1175342430 CA339961314 |
103 | A>D | No |
ClinGen TOPMed |
|
|
rs755330447 CA803563 |
103 | A>S | No |
ClinGen ExAC gnomAD |
|
|
rs1570593595 RCV000993013 |
106 | S>missing | No |
ClinVar dbSNP |
|
|
rs775307302 COSM167057 CA803557 |
107 | A>T | large_intestine [Cosmic] | No |
ClinGen cosmic curated ExAC gnomAD |
|
rs1377462340 CA339961250 |
107 | A>V | No |
ClinGen Ensembl |
|
|
CA339961231 rs1436569888 |
109 | L>F | No |
ClinGen gnomAD |
|
| TCGA novel | 111 | G>C | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
CA803551 rs769786931 |
116 | G>D | No |
ClinGen ExAC gnomAD |
|
|
rs777945822 CA803552 |
116 | G>S | No |
ClinGen ExAC gnomAD |
|
|
CA803550 rs748648403 |
117 | K>R | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA339961024 rs1172279073 |
122 | L>V | No |
ClinGen gnomAD |
|
|
CA803549 rs781521534 |
125 | G>A | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA318427 RCV000189352 rs80359819 |
130 | G>R | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs113226716 CA21252045 |
132 | Y>H | No |
ClinGen Ensembl |
|
|
rs1553156161 CA658656902 RCV000519895 |
133 | C>F | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs1570593487 RCV000993978 |
137 | T>missing | No |
ClinVar dbSNP |
|
|
rs780519584 CA339960746 |
142 | M>L | No |
ClinGen gnomAD |
|
|
rs780519584 CA21252033 |
142 | M>V | No |
ClinGen gnomAD |
|
|
CA803544 rs760130033 |
147 | V>L | No |
ClinGen ExAC gnomAD |
|
|
CA339960612 rs1277225461 |
149 | P>H | No |
ClinGen gnomAD |
|
|
rs752289598 CA803543 |
150 | T>A | No |
ClinGen ExAC gnomAD |
|
|
CA339960577 rs1351005021 |
151 | A>D | No |
ClinGen gnomAD |
|
| TCGA novel | 157 | G>D | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
RCV000503688 CA339960486 rs1553156155 |
158 | T>P | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
CA339960436 RCV000512938 rs1553156154 |
161 | Q>R | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs1085307483 RCV000489368 CA339960405 |
163 | G>S | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs770867924 CA803539 |
166 | V>I | No |
ClinGen ExAC TOPMed gnomAD |
|
|
rs773339124 CA803538 |
167 | G>S | No |
ClinGen ExAC gnomAD |
|
|
RCV000485695 rs1553156146 |
172 | Q>missing | No |
ClinVar dbSNP |
|
|
rs1570592968 CA339958957 |
173 | V>G | No |
ClinGen Ensembl |
|
|
CA339958848 rs1570592958 |
179 | I>V | No |
ClinGen Ensembl |
|
|
RCV000189386 rs796053267 |
180 | M>missing | No |
ClinVar dbSNP |
|
|
CA339958750 rs1557646134 |
183 | K>E | No |
ClinGen Ensembl |
|
|
CA803497 rs746393667 |
192 | I>L | No |
ClinGen ExAC gnomAD |
|
|
rs1212331558 CA339958663 |
193 | I>V | No |
ClinGen gnomAD |
|
| TCGA novel | 193 | I>F | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
rs368782332 CA21251480 |
193 | I>T | No |
ClinGen Ensembl |
|
|
CA803496 rs774524003 |
195 | I>F | No |
ClinGen ExAC gnomAD |
|
|
CA803495 rs771352226 |
196 | P>L | No |
ClinGen ExAC gnomAD |
|
|
rs1381362306 CA339958634 |
197 | A>V | No |
ClinGen TOPMed |
|
|
rs1324984353 CA339958621 |
200 | Q>R | No |
ClinGen TOPMed |
|
|
CA21251447 rs921770589 |
201 | C>F | No |
ClinGen Ensembl |
|
|
CA803490 rs758200582 |
203 | V>L | No |
ClinGen ExAC gnomAD |
|
|
rs1402428875 CA339958594 |
205 | P>A | No |
ClinGen gnomAD |
|
|
rs1304291329 CA339958587 |
206 | F>L | No |
ClinGen gnomAD |
|
| TCGA novel | 208 | P>L | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
rs1159263312 CA339958534 |
213 | F>L | No |
ClinGen gnomAD |
|
|
rs374080633 CA803487 |
218 | R>L | No |
ClinGen ESP ExAC TOPMed gnomAD |
|
|
CA339958492 rs1257108581 |
220 | E>A | No |
ClinGen gnomAD |
|
|
CA339958496 rs1448464718 |
220 | E>K | Variant assessed as Somatic; impact. [NCI-TCGA] | No |
ClinGen NCI-TCGA TOPMed |
| TCGA novel | 221 | E>R | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
CA21251395 rs934104576 |
222 | N>S | No |
ClinGen TOPMed |
|
|
rs767600686 CA803483 |
224 | A>T | No |
ClinGen ExAC gnomAD |
|
|
CA339958460 rs1209468941 |
225 | K>N | No |
ClinGen gnomAD |
|
|
CA339958455 rs1261396902 |
226 | S>R | No |
ClinGen TOPMed gnomAD |
|
|
rs1570592655 RCV001009153 |
233 | G>missing | No |
ClinVar dbSNP |
|
|
CA339958210 rs1466673582 |
235 | A>V | No |
ClinGen gnomAD |
|
|
rs796053249 CA318434 RCV000189357 |
236 | D>V | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs771662171 CA803461 |
236 | D>Y | No |
ClinGen ExAC gnomAD |
|
|
CA339958183 rs1387242348 |
237 | V>M | Variant assessed as Somatic; 4.621e-05 impact. [NCI-TCGA] | No |
ClinGen NCI-TCGA TOPMed gnomAD |
|
rs1045642109 CA21251163 |
239 | H>R | No |
ClinGen TOPMed gnomAD |
|
|
CA339958080 rs1305932942 |
244 | M>L | No |
ClinGen gnomAD |
|
|
CA16617146 rs1064795363 RCV000482535 |
244 | M>R | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs773788706 CA803460 |
245 | K>R | No |
ClinGen ExAC gnomAD |
|
|
CA21251131 rs80359821 RCV000494114 |
247 | E>D | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs749226083 CA803459 |
251 | M>T | No |
ClinGen ExAC TOPMed gnomAD |
|
| TCGA novel | 251 | M>V | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
rs1013744464 CA21251114 |
252 | M>V | No |
ClinGen Ensembl |
|
|
CA339957925 rs1451575059 |
253 | R>G | No |
ClinGen TOPMed gnomAD |
|
|
CA339957918 rs777498634 |
253 | R>P | No |
ClinGen ExAC TOPMed gnomAD |
|
|
rs777498634 CA803458 |
253 | R>Q | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA339957924 rs1451575059 |
253 | R>W | No |
ClinGen TOPMed gnomAD |
|
|
rs1184570993 CA339957883 |
255 | K>N | No |
ClinGen gnomAD |
|
|
rs5811 CA803457 |
255 | K>R | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA21251076 rs935400435 |
256 | K>M | No |
ClinGen TOPMed gnomAD |
|
|
CA803456 rs781458890 |
256 | K>N | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA339957875 rs935400435 |
256 | K>T | No |
ClinGen TOPMed gnomAD |
|
|
CA803455 rs755213963 |
257 | V>F | No |
ClinGen ExAC |
|
|
rs1570592543 CA339957853 |
257 | V>G | No |
ClinGen Ensembl |
|
|
rs200221625 CA21251033 |
265 | S>P | No |
ClinGen Ensembl |
|
|
rs199639208 CA339957599 |
267 | A>S | No |
ClinGen 1000Genomes ESP ExAC TOPMed gnomAD |
|
|
rs121909740 CA339957376 |
275 | A>S | No |
ClinGen gnomAD |
|
|
rs145079633 CA21250993 |
276 | V>M | No |
ClinGen ESP |
|
|
RCV000189387 rs796053268 |
277 | V>missing | No |
ClinVar dbSNP |
|
|
CA21250989 rs928228807 |
277 | V>L | No |
ClinGen TOPMed |
|
|
rs796053251 CA318440 RCV000189362 |
283 | Q>R | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
CA339957164 rs1431778557 |
287 | I>V | No |
ClinGen TOPMed |
|
| TCGA novel | 289 | A>= | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
RCV000189363 RCV002252036 CA318442 rs796053252 |
289 | A>T | No |
ClinGen ClinVar TOPMed dbSNP |
|
|
CA339956973 rs1570592186 |
290 | V>G | No |
ClinGen Ensembl |
|
|
rs753156126 CA803408 |
290 | V>I | No |
ClinGen ExAC gnomAD |
|
|
RCV000483051 rs1064795152 |
292 | Y>missing | No |
ClinVar dbSNP |
|
| TCGA novel | 296 | S>G | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
rs761764013 CA803405 |
300 | K>N | No |
ClinGen ExAC gnomAD |
|
|
CA803406 rs764743728 |
300 | K>T | No |
ClinGen ExAC gnomAD |
|
|
CA339956774 rs944584284 |
302 | G>A | No |
ClinGen gnomAD |
|
|
CA21250704 rs944584284 |
302 | G>E | No |
ClinGen gnomAD |
|
|
CA339956777 rs1228422579 |
302 | G>W | No |
ClinGen TOPMed |
|
|
rs80359837 RCV000179917 RCV000478834 |
303 | V>missing | No |
ClinVar dbSNP |
|
|
CA339956746 rs1570592136 |
303 | V>G | No |
ClinGen Ensembl |
|
|
rs1253340181 CA339956681 |
307 | V>L | No |
ClinGen TOPMed gnomAD |
|
|
rs1253340181 CA339956685 |
307 | V>M | No |
ClinGen TOPMed gnomAD |
|
|
rs749844687 CA21250639 |
311 | I>T | No |
ClinGen Ensembl |
|
|
RCV000329396 rs886041633 CA10602779 |
313 | S>P | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
CA019355 rs794727870 RCV000179922 |
313 | S>Y | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs1349205424 CA339956322 |
322 | V>L | No |
ClinGen gnomAD |
|
|
rs886044287 RCV000341326 |
324 | S>missing | No |
ClinVar dbSNP |
|
|
RCV000520273 rs1553156002 CA339956285 |
324 | S>P | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs747411639 CA339956146 |
326 | F>C | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA803374 rs747411639 |
326 | F>S | No |
ClinGen ExAC TOPMed gnomAD |
|
|
rs796053270 RCV000189390 |
327 | V>missing | No |
ClinVar dbSNP |
|
|
rs80359838 RCV000180298 |
327 | V>missing | No |
ClinVar dbSNP |
|
|
rs796053255 CA318451 RCV000189368 |
328 | V>L | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs80359826 COSM3689926 CA318438 |
330 | R>* | large_intestine Variant assessed as Somatic; impact. [Cosmic, NCI-TCGA] | No |
ClinGen cosmic curated ExAC NCI-TCGA gnomAD |
|
rs753278619 CA803371 |
330 | R>Q | No |
ClinGen ExAC gnomAD |
|
|
CA339956049 rs1225524483 |
331 | A>V | No |
ClinGen gnomAD |
|
|
rs1363752047 CA339956030 |
332 | G>D | No |
ClinGen gnomAD |
|
| TCGA novel | 334 | R>W | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
CA339955979 rs1318443924 |
335 | T>I | No |
ClinGen gnomAD |
|
|
RCV000189391 rs796053271 |
336 | L>missing | No |
ClinVar dbSNP |
|
|
CA803369 rs755946345 |
336 | L>M | No |
ClinGen ExAC gnomAD |
|
|
rs2229681 CA339955931 |
337 | H>Q | No |
ClinGen 1000Genomes ESP ExAC TOPMed gnomAD |
|
|
rs752703331 CA803368 |
337 | H>Y | No |
ClinGen ExAC gnomAD |
|
|
CA339955918 rs1386083305 |
338 | L>F | No |
ClinGen gnomAD |
|
|
CA318453 RCV000189369 rs796053256 |
339 | I>V | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs886041981 RCV000372651 |
340 | G>missing | No |
ClinVar dbSNP |
|
| TCGA novel | 340 | G>S | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
rs1430786115 CA339955783 |
344 | M>I | No |
ClinGen gnomAD |
|
|
rs769197923 CA803363 |
348 | A>V | No |
ClinGen ExAC gnomAD |
|
|
CA339955685 rs1202505639 |
349 | I>L | No |
ClinGen TOPMed gnomAD |
|
|
rs1341049777 CA339955663 |
349 | I>T | No |
ClinGen gnomAD |
|
|
CA339955678 rs1202505639 |
349 | I>V | No |
ClinGen TOPMed gnomAD |
|
|
CA318455 rs796053257 RCV000189371 |
350 | L>P | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs1270248389 CA339955606 |
351 | M>I | No |
ClinGen gnomAD |
|
| TCGA novel | 352 | T>A | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
CA21250132 rs531005993 |
352 | T>I | No |
ClinGen 1000Genomes |
|
|
CA21250128 rs992419688 |
353 | I>T | No |
ClinGen Ensembl |
|
|
rs1229028572 CA339955574 |
353 | I>V | No |
ClinGen gnomAD |
|
|
CA803360 rs756790359 |
354 | A>V | No |
ClinGen ExAC gnomAD |
|
|
CA339955483 rs1454569723 |
357 | L>P | No |
ClinGen TOPMed gnomAD |
|
|
RCV001008987 rs1570591037 |
359 | E>missing | No |
ClinVar dbSNP |
|
|
rs1553155900 RCV000598861 |
362 | P>missing | No |
ClinVar dbSNP |
|
|
rs747658630 CA803338 |
362 | P>S | No |
ClinGen ExAC TOPMed gnomAD |
|
|
rs1203803892 CA339954407 |
364 | M>L | No |
ClinGen gnomAD |
|
|
rs1325549402 CA339954396 |
364 | M>T | No |
ClinGen gnomAD |
|
|
CA339954322 rs1395523942 |
368 | S>N | No |
ClinGen Ensembl |
|
|
CA339954299 rs1227537819 |
369 | I>N | No |
ClinGen gnomAD |
|
|
CA318457 RCV000189372 rs796053258 |
369 | I>V | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
CA803335 rs779678519 |
377 | A>T | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA21249189 rs75020603 |
380 | E>* | No |
ClinGen Ensembl |
|
|
rs929498798 CA21249187 |
381 | V>M | No |
ClinGen Ensembl |
|
|
CA16603688 rs1057521632 RCV000442546 |
383 | P>L | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs886039737 RCV000255483 |
386 | I>missing | No |
ClinVar dbSNP |
|
|
rs1553155889 CA318465 |
386 | I>V | No |
ClinGen Ensembl |
|
|
CA339953916 rs1384263716 |
392 | A>P | No |
ClinGen gnomAD |
|
|
rs1570590940 CA339953904 |
393 | E>D | No |
ClinGen Ensembl |
|
|
CA658656901 rs1553155885 RCV000520991 |
396 | S>* | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
CA339953887 rs1183662521 |
396 | S>G | No |
ClinGen gnomAD |
|
|
CA21249167 rs80359827 |
397 | Q>* | No |
ClinGen Ensembl |
|
|
rs1027810691 CA21249165 |
398 | G>S | No |
ClinGen TOPMed gnomAD |
|
|
rs1557645029 CA339953827 |
402 | A>P | No |
ClinGen Ensembl |
|
|
CA803325 rs774646485 |
404 | I>T | No |
ClinGen ExAC gnomAD |
|
|
rs374789687 CA803326 |
404 | I>V | No |
ClinGen ESP ExAC TOPMed gnomAD |
|
|
rs1264080435 CA339953744 |
407 | A>G | No |
ClinGen gnomAD |
|
|
rs1318652576 CA339953722 |
409 | F>V | No |
ClinGen gnomAD |
|
|
RCV000599180 rs1553155880 |
410 | S>missing | No |
ClinVar dbSNP |
|
| TCGA novel | 418 | V>A | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
CA21249071 rs139722450 |
419 | G>A | No |
ClinGen ESP TOPMed |
|
|
rs1320662318 CA339953653 |
419 | G>S | No |
ClinGen gnomAD |
|
|
RCV000413602 CA16042369 rs139722450 |
419 | G>V | No |
ClinGen ClinVar ESP TOPMed dbSNP |
|
|
CA339953619 rs1425712703 |
423 | Q>H | No |
ClinGen TOPMed |
|
|
RCV000147521 rs587784392 CA019033 |
423 | Q>R | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
rs1188094356 CA339953607 |
425 | V>A | No |
ClinGen Ensembl |
|
|
rs1553155843 RCV000522544 CA339953583 |
427 | Q>* | No |
ClinGen ClinVar Ensembl dbSNP |
|
|
CA339953533 rs1175838250 |
431 | P>L | No |
ClinGen gnomAD |
|
| TCGA novel | 431 | P>S | Variant assessed as Somatic; impact. [NCI-TCGA] | No | NCI-TCGA |
|
rs1468239807 CA339953528 |
432 | Y>H | No |
ClinGen gnomAD |
|
|
rs200819771 CA803289 |
433 | V>I | No |
ClinGen 1000Genomes ExAC TOPMed gnomAD |
|
|
CA803287 rs756347604 |
449 | Y>F | No |
ClinGen ExAC gnomAD |
|
|
rs1216296247 CA339953247 |
450 | F>L | No |
ClinGen TOPMed |
|
|
CA21248814 rs767510220 |
457 | G>A | No |
ClinGen ExAC gnomAD |
|
|
CA803285 rs767510220 |
457 | G>D | No |
ClinGen ExAC gnomAD |
|
|
rs752143706 RCV000440623 CA803284 |
458 | R>Q | No |
ClinGen ClinVar ExAC dbSNP gnomAD |
|
|
rs1243627480 CA339952993 |
461 | D>E | No |
ClinGen TOPMed |
|
|
CA21248791 rs370257930 |
461 | D>N | No |
ClinGen ESP |
|
|
CA21248787 rs1047721769 |
461 | D>V | No |
ClinGen TOPMed |
|
|
rs763390868 CA803282 |
465 | S>T | No |
ClinGen ExAC gnomAD |
|
|
rs1225304808 CA339952887 |
466 | G>D | Variant assessed as Somatic; impact. [NCI-TCGA] | No |
ClinGen NCI-TCGA TOPMed |
|
rs13306754 CA21248775 |
468 | R>L | No |
ClinGen gnomAD |
|
|
CA339952848 rs13306754 |
468 | R>Q | No |
ClinGen gnomAD |
|
|
rs1553155816 RCV000598793 |
469 | Q>missing | No |
ClinVar dbSNP |
|
|
rs1421901500 CA339952726 |
473 | S>G | No |
ClinGen TOPMed |
|
|
CA21248747 rs994163843 |
478 | T>I | No |
ClinGen Ensembl |
|
|
rs1025285774 CA21248750 |
478 | T>P | No |
ClinGen Ensembl |
|
|
CA339952415 rs1362971897 |
483 | F>S | No |
ClinGen gnomAD |
|
|
rs1159593580 CA339952342 |
485 | P>R | No |
ClinGen gnomAD |
|
|
rs1460882564 CA339952202 |
490 | S>T | No |
ClinGen gnomAD |
|
|
CA339952165 rs754985269 |
492 | V>L | No |
ClinGen ExAC TOPMed gnomAD |
|
|
CA803275 rs754985269 |
492 | V>M | No |
ClinGen ExAC TOPMed gnomAD |
5 associated diseases with P11166
[MIM: 606777]: GLUT1 deficiency syndrome 1 (GLUT1DS1)
A neurologic disorder showing wide phenotypic variability. The most severe 'classic' phenotype comprises infantile-onset epileptic encephalopathy associated with delayed development, acquired microcephaly, motor incoordination, and spasticity. Onset of seizures, usually characterized by apneic episodes, staring spells, and episodic eye movements, occurs within the first 4 months of life. Other paroxysmal findings include intermittent ataxia, confusion, lethargy, sleep disturbance, and headache. Varying degrees of cognitive impairment can occur, ranging from learning disabilities to severe intellectual disability. {ECO:0000269|PubMed:10227690, ECO:0000269|PubMed:10980529, ECO:0000269|PubMed:11136715, ECO:0000269|PubMed:11603379, ECO:0000269|PubMed:12325075, ECO:0000269|PubMed:15622525, ECO:0000269|PubMed:19901175, ECO:0000269|PubMed:20129935, ECO:0000269|PubMed:20221955, ECO:0000269|PubMed:20574033, ECO:0000269|PubMed:24847886, ECO:0000269|PubMed:25982116, ECO:0000269|PubMed:30197081}. Note=The disease is caused by variants affecting the gene represented in this entry.
[MIM: 612126]: GLUT1 deficiency syndrome 2 (GLUT1DS2)
A clinically variable disorder characterized primarily by onset in childhood of paroxysmal exercise-induced dyskinesia. The dyskinesia involves transient abnormal involuntary movements, such as dystonia and choreoathetosis, induced by exercise or exertion, and affecting the exercised limbs. Some patients may also have epilepsy, most commonly childhood absence epilepsy. Mild intellectual disability may also occur. In some patients involuntary exertion-induced dystonic, choreoathetotic, and ballistic movements may be associated with macrocytic hemolytic anemia. {ECO:0000269|PubMed:14605501, ECO:0000269|PubMed:18451999, ECO:0000269|PubMed:19630075, ECO:0000269|PubMed:19798636, ECO:0000269|PubMed:20129935, ECO:0000269|PubMed:20574033, ECO:0000269|PubMed:20621801, ECO:0000269|PubMed:20830593, ECO:0000269|PubMed:21204808}. Note=The disease is caused by variants affecting the gene represented in this entry.
[MIM: 614847]: Epilepsy, idiopathic generalized 12 (EIG12)
A disorder characterized by recurring generalized seizures in the absence of detectable brain lesions and/or metabolic abnormalities. Generalized seizures arise diffusely and simultaneously from both hemispheres of the brain. Seizure types include juvenile myoclonic seizures, absence seizures, and generalized tonic-clonic seizures. In some EIG12 patients seizures may remit with age. {ECO:0000269|PubMed:19798636, ECO:0000269|PubMed:22282645, ECO:0000269|PubMed:23280796, ECO:0000269|PubMed:25982116}. Note=Disease susceptibility is associated with variants affecting the gene represented in this entry.
[MIM: 601042]: Dystonia 9 (DYT9)
An autosomal dominant neurologic disorder characterized by childhood onset of paroxysmal choreoathetosis and progressive spastic paraplegia. Most patients show some degree of cognitive impairment. Other variable features may include seizures, migraine headaches, and ataxia. {ECO:0000269|PubMed:21832227}. Note=The disease is caused by variants affecting the gene represented in this entry.
[MIM: 608885]: Stomatin-deficient cryohydrocytosis with neurologic defects (SDCHCN)
A rare form of stomatocytosis characterized by episodic hemolytic anemia, cold-induced red cells cation leak, erratic hyperkalemia, neonatal hyperbilirubinemia, hepatosplenomegaly, cataracts, seizures, intellectual disability, and movement disorder. {ECO:0000269|PubMed:21791420, ECO:0000269|PubMed:22492876}. Note=The disease is caused by variants affecting the gene represented in this entry.
Without disease ID
- A neurologic disorder showing wide phenotypic variability. The most severe 'classic' phenotype comprises infantile-onset epileptic encephalopathy associated with delayed development, acquired microcephaly, motor incoordination, and spasticity. Onset of seizures, usually characterized by apneic episodes, staring spells, and episodic eye movements, occurs within the first 4 months of life. Other paroxysmal findings include intermittent ataxia, confusion, lethargy, sleep disturbance, and headache. Varying degrees of cognitive impairment can occur, ranging from learning disabilities to severe intellectual disability. {ECO:0000269|PubMed:10227690, ECO:0000269|PubMed:10980529, ECO:0000269|PubMed:11136715, ECO:0000269|PubMed:11603379, ECO:0000269|PubMed:12325075, ECO:0000269|PubMed:15622525, ECO:0000269|PubMed:19901175, ECO:0000269|PubMed:20129935, ECO:0000269|PubMed:20221955, ECO:0000269|PubMed:20574033, ECO:0000269|PubMed:24847886, ECO:0000269|PubMed:25982116, ECO:0000269|PubMed:30197081}. Note=The disease is caused by variants affecting the gene represented in this entry.
- A clinically variable disorder characterized primarily by onset in childhood of paroxysmal exercise-induced dyskinesia. The dyskinesia involves transient abnormal involuntary movements, such as dystonia and choreoathetosis, induced by exercise or exertion, and affecting the exercised limbs. Some patients may also have epilepsy, most commonly childhood absence epilepsy. Mild intellectual disability may also occur. In some patients involuntary exertion-induced dystonic, choreoathetotic, and ballistic movements may be associated with macrocytic hemolytic anemia. {ECO:0000269|PubMed:14605501, ECO:0000269|PubMed:18451999, ECO:0000269|PubMed:19630075, ECO:0000269|PubMed:19798636, ECO:0000269|PubMed:20129935, ECO:0000269|PubMed:20574033, ECO:0000269|PubMed:20621801, ECO:0000269|PubMed:20830593, ECO:0000269|PubMed:21204808}. Note=The disease is caused by variants affecting the gene represented in this entry.
- A disorder characterized by recurring generalized seizures in the absence of detectable brain lesions and/or metabolic abnormalities. Generalized seizures arise diffusely and simultaneously from both hemispheres of the brain. Seizure types include juvenile myoclonic seizures, absence seizures, and generalized tonic-clonic seizures. In some EIG12 patients seizures may remit with age. {ECO:0000269|PubMed:19798636, ECO:0000269|PubMed:22282645, ECO:0000269|PubMed:23280796, ECO:0000269|PubMed:25982116}. Note=Disease susceptibility is associated with variants affecting the gene represented in this entry.
- An autosomal dominant neurologic disorder characterized by childhood onset of paroxysmal choreoathetosis and progressive spastic paraplegia. Most patients show some degree of cognitive impairment. Other variable features may include seizures, migraine headaches, and ataxia. {ECO:0000269|PubMed:21832227}. Note=The disease is caused by variants affecting the gene represented in this entry.
- A rare form of stomatocytosis characterized by episodic hemolytic anemia, cold-induced red cells cation leak, erratic hyperkalemia, neonatal hyperbilirubinemia, hepatosplenomegaly, cataracts, seizures, intellectual disability, and movement disorder. {ECO:0000269|PubMed:21791420, ECO:0000269|PubMed:22492876}. Note=The disease is caused by variants affecting the gene represented in this entry.
3 regional properties for P11166
| Type | Name | Position | InterPro Accession |
|---|---|---|---|
| conserved_site | Sugar transporter, conserved site | 128 - 153 | IPR005829-1 |
| conserved_site | Sugar transporter, conserved site | 324 - 340 | IPR005829-2 |
| domain | Major facilitator superfamily domain | 15 - 456 | IPR020846 |
Functions
20 GO annotations of cellular component
| Name | Definition |
|---|---|
| apical plasma membrane | The region of the plasma membrane located at the apical end of the cell. |
| basolateral plasma membrane | The region of the plasma membrane that includes the basal end and sides of the cell. Often used in reference to animal polarized epithelial membranes, where the basal membrane is the part attached to the extracellular matrix, or in plant cells, where the basal membrane is defined with respect to the zygotic axis. |
| blood microparticle | A phospholipid microvesicle that is derived from any of several cell types, such as platelets, blood cells, endothelial cells, or others, and contains membrane receptors as well as other proteins characteristic of the parental cell. Microparticles are heterogeneous in size, and are characterized as microvesicles free of nucleic acids. |
| caveola | A membrane raft that forms small pit, depression, or invagination that communicates with the outside of a cell and extends inward, indenting the cytoplasm and the cell membrane. Examples include flask-shaped invaginations of the plasma membrane in adipocytes associated with caveolin proteins, and minute pits or incuppings of the cell membrane formed during pinocytosis. Caveolae may be pinched off to form free vesicles within the cytoplasm. |
| cortical actin cytoskeleton | The portion of the actin cytoskeleton, comprising filamentous actin and associated proteins, that lies just beneath the plasma membrane. |
| cytosol | The part of the cytoplasm that does not contain organelles but which does contain other particulate matter, such as protein complexes. |
| extracellular exosome | A vesicle that is released into the extracellular region by fusion of the limiting endosomal membrane of a multivesicular body with the plasma membrane. Extracellular exosomes, also simply called exosomes, have a diameter of about 40-100 nm. |
| female germ cell nucleus | The nucleus of the female germ cell, a reproductive cell in females. |
| female pronucleus | The pronucleus originating from the ovum that is being fertilized. |
| Golgi membrane | The lipid bilayer surrounding any of the compartments of the Golgi apparatus. |
| integral component of plasma membrane | The component of the plasma membrane consisting of the gene products and protein complexes having at least some part of their peptide sequence embedded in the hydrophobic region of the membrane. |
| intercalated disc | A complex cell-cell junction at which myofibrils terminate in cardiomyocytes; mediates mechanical and electrochemical integration between individual cardiomyocytes. The intercalated disc contains regions of tight mechanical attachment (fasciae adherentes and desmosomes) and electrical coupling (gap junctions) between adjacent cells. |
| melanosome | A tissue-specific, membrane-bounded cytoplasmic organelle within which melanin pigments are synthesized and stored. Melanosomes are synthesized in melanocyte cells. |
| membrane | A lipid bilayer along with all the proteins and protein complexes embedded in it an attached to it. |
| midbody | A thin cytoplasmic bridge formed between daughter cells at the end of cytokinesis. The midbody forms where the contractile ring constricts, and may persist for some time before finally breaking to complete cytokinesis. |
| photoreceptor inner segment | The inner segment of a vertebrate photoreceptor containing mitochondria, ribosomes and membranes where opsin molecules are assembled and passed to be part of the outer segment discs. |
| plasma membrane | The membrane surrounding a cell that separates the cell from its external environment. It consists of a phospholipid bilayer and associated proteins. |
| presynapse | The part of a synapse that is part of the presynaptic cell. |
| sarcolemma | The outer membrane of a muscle cell, consisting of the plasma membrane, a covering basement membrane (about 100 nm thick and sometimes common to more than one fiber), and the associated loose network of collagen fibers. |
| Z disc | Platelike region of a muscle sarcomere to which the plus ends of actin filaments are attached. |
9 GO annotations of molecular function
| Name | Definition |
|---|---|
| D-glucose transmembrane transporter activity | Enables the transfer of the D-enantiomer of the hexose monosaccharide glucose from one side of a membrane to the other. |
| dehydroascorbic acid transmembrane transporter activity | Enables the transfer of dehydroascorbate, 5-(1,2-dihydroxyethyl)furan-2,3,4(5H)-trione, from one side of a membrane to the other. |
| glucose transmembrane transporter activity | Enables the transfer of the hexose monosaccharide glucose from one side of a membrane to the other. |
| hexose transmembrane transporter activity | Enables the transfer of a hexose sugar, a monosaccharide with 6 carbon atoms, from one side of a membrane to the other. |
| identical protein binding | Binding to an identical protein or proteins. |
| kinase binding | Binding to a kinase, any enzyme that catalyzes the transfer of a phosphate group. |
| long-chain fatty acid transporter activity | Enables the transfer of long-chain fatty acids from one side of a membrane to the other. A long-chain fatty acid is a fatty acid with a chain length between C13 and C22. |
| protein self-association | Binding to a domain within the same polypeptide. |
| xenobiotic transmembrane transporter activity | Enables the directed movement of a xenobiotic from one side of a membrane to the other. A xenobiotic is a compound foreign to the organim exposed to it. It may be synthesized by another organism (like ampicilin) or it can be a synthetic chemical. |
19 GO annotations of biological process
| Name | Definition |
|---|---|
| cellular hyperosmotic response | Any process that results in a change in state or activity of a cell (in terms of movement, secretion, enzyme production, gene expression, etc.) as a result of detection of, or exposure to, a hyperosmotic environment, i.e. an environment with a higher concentration of solutes than the organism or cell. |
| cellular response to glucose starvation | Any process that results in a change in state or activity of a cell (in terms of movement, secretion, enzyme production, gene expression, etc.) as a result of deprivation of glucose. |
| cellular response to mechanical stimulus | Any process that results in a change in state or activity of a cell (in terms of movement, secretion, enzyme production, gene expression, etc.) as a result of a mechanical stimulus. |
| central nervous system development | The process whose specific outcome is the progression of the central nervous system over time, from its formation to the mature structure. The central nervous system is the core nervous system that serves an integrating and coordinating function. In vertebrates it consists of the brain and spinal cord. In those invertebrates with a central nervous system it typically consists of a brain, cerebral ganglia and a nerve cord. |
| cerebral cortex development | The progression of the cerebral cortex over time from its initial formation until its mature state. The cerebral cortex is the outer layered region of the telencephalon. |
| dehydroascorbic acid transport | The directed movement of dehydroascorbate into, out of or within a cell, or between cells, by means of some agent such as a transporter or pore. Dehydroascorbate, 5-(1,2-dihydroxyethyl)furan-2,3,4(5H)-trione, is an oxidized form of vitamin C. |
| female pregnancy | The set of physiological processes that allow an embryo or foetus to develop within the body of a female animal. It covers the time from fertilization of a female ovum by a male spermatozoon until birth. |
| glucose import | The directed movement of the hexose monosaccharide glucose into a cell or organelle. |
| glucose import across plasma membrane | The directed movement of glucose from outside of a cell, across the plasma membrane and into the cytosol. |
| glucose transmembrane transport | The process in which glucose is transported across a membrane. |
| L-ascorbic acid metabolic process | The chemical reactions and pathways involving L-ascorbic acid, (2R)-2-[(1S)-1,2-dihydroxyethyl]-4-hydroxy-5-oxo-2,5-dihydrofuran-3-olate; L-ascorbic acid is vitamin C and has co-factor and anti-oxidant activities in many species. |
| long-chain fatty acid import across plasma membrane | The directed movement of long-chain fatty acids from outside of a cell, across the plasma membrane and into the cytosol. A long-chain fatty acid is a fatty acid with a chain length between C13 and C22. |
| monosaccharide transmembrane transport | The process in which a monosaccharide is transported across a lipid bilayer, from one side of a membrane to the other. Monosaccharides are the simplest carbohydrates; they are polyhydric alcohols containing either an aldehyde or a keto group and between three to ten or more carbon atoms. They form the constitutional repeating units of oligo- and polysaccharides. |
| photoreceptor cell maintenance | Any process preventing the degeneration of the photoreceptor, a specialized cell type that is sensitive to light. |
| protein-containing complex assembly | The aggregation, arrangement and bonding together of a set of macromolecules to form a protein-containing complex. |
| response to hypoxia | Any process that results in a change in state or activity of a cell or an organism (in terms of movement, secretion, enzyme production, gene expression, etc.) as a result of a stimulus indicating lowered oxygen tension. Hypoxia, defined as a decline in O2 levels below normoxic levels of 20.8 - 20.95%, results in metabolic adaptation at both the cellular and organismal level. |
| response to insulin | Any process that results in a change in state or activity of a cell or an organism (in terms of movement, secretion, enzyme production, gene expression, etc.) as a result of an insulin stimulus. Insulin is a polypeptide hormone produced by the islets of Langerhans of the pancreas in mammals, and by the homologous organs of other organisms. |
| response to Thyroglobulin triiodothyronine | Any process that results in a change in state or activity of a cell or an organism (in terms of movement, secretion, enzyme production, gene expression, etc.) as a result of a Thyroglobulin triiodothyronine stimulus. |
| transport across blood-brain barrier | The directed movement of substances (e.g. macromolecules, small molecules, ions) through the blood-brain barrier. |
4 homologous proteins in AiPD
| UniProt AC | Gene Name | Protein Name | Species | Evidence Code |
|---|---|---|---|---|
| P58352 | SLC2A3 | Solute carrier family 2, facilitated glucose transporter member 3 | Bos taurus (Bovine) | PR |
| P27674 | SLC2A1 | Solute carrier family 2, facilitated glucose transporter member 1 | Bos taurus (Bovine) | PR |
| P17809 | Slc2a1 | Solute carrier family 2, facilitated glucose transporter member 1 | Mus musculus (Mouse) | PR |
| P20303 | SLC2A1 | Solute carrier family 2, facilitated glucose transporter member 1 | Sus scrofa (Pig) | PR |
| 10 | 20 | 30 | 40 | 50 | 60 |
| MEPSSKKLTG | RLMLAVGGAV | LGSLQFGYNT | GVINAPQKVI | EEFYNQTWVH | RYGESILPTT |
| 70 | 80 | 90 | 100 | 110 | 120 |
| LTTLWSLSVA | IFSVGGMIGS | FSVGLFVNRF | GRRNSMLMMN | LLAFVSAVLM | GFSKLGKSFE |
| 130 | 140 | 150 | 160 | 170 | 180 |
| MLILGRFIIG | VYCGLTTGFV | PMYVGEVSPT | ALRGALGTLH | QLGIVVGILI | AQVFGLDSIM |
| 190 | 200 | 210 | 220 | 230 | 240 |
| GNKDLWPLLL | SIIFIPALLQ | CIVLPFCPES | PRFLLINRNE | ENRAKSVLKK | LRGTADVTHD |
| 250 | 260 | 270 | 280 | 290 | 300 |
| LQEMKEESRQ | MMREKKVTIL | ELFRSPAYRQ | PILIAVVLQL | SQQLSGINAV | FYYSTSIFEK |
| 310 | 320 | 330 | 340 | 350 | 360 |
| AGVQQPVYAT | IGSGIVNTAF | TVVSLFVVER | AGRRTLHLIG | LAGMAGCAIL | MTIALALLEQ |
| 370 | 380 | 390 | 400 | 410 | 420 |
| LPWMSYLSIV | AIFGFVAFFE | VGPGPIPWFI | VAELFSQGPR | PAAIAVAGFS | NWTSNFIVGM |
| 430 | 440 | 450 | 460 | 470 | 480 |
| CFQYVEQLCG | PYVFIIFTVL | LVLFFIFTYF | KVPETKGRTF | DEIASGFRQG | GASQSDKTPE |
| 490 | |||||
| ELFHPLGADS | QV |